Today was the end of NMR and we discussed multiplicity. We say the theoretical basis of the "N+1 Rule" and looked at a bunch of examples. The main point was essentially this: The integration is an indicator of how many hydrogens are responsible for the resonance and the multiplicity is an indication of the number of hydrogens on adjacent carbon atoms. We saw singlets, doublets, triplets and quartets. Complicated cases, where two or more different (non-equivalent) sets of hydrogens were nearby, led to multiplets.
We looked at the problem set and the 11 pages of the problem set key and saw the on-line problem set from Notre Dame (link at left). Each group, in lieu of a mash-up, is to give me the answers to the 12 listed problems by next Friday. Help each other!
Friday, April 13, 2007

Thursday, April 12, 2007
1H NMR
Yesterday was a continuation of NMR, except we looked at the NMR of hydrogen rather than carbon. The concepts are the same, with an exception or two.....the main difference that we saw was that there is a quantitative aspect to 1H NMR that is not present in 13C NMR. It is possible to do an integration of the peaks, which will give us a relative ratio of the number of hydrogens responsible for each signal. Yesterday, all of the peaks were single peaks, on Friday we will look at how peaks can have different multiplicities -- there was some foreshadowing when I mentioned the N+1 Rule at the end of class.
Many changes have been made to the left: The Need to Knows for Chapters 13, 14 and 15 have been posted, as have the charts for 13C and 1H NMR. (Remember, you'll be given the 13C chart but must learn the 1H chart.) Modeling exercise #10 has also been posted; I'll talk more about it tomorrow.
New changes! A key for the exam and an NMR problem set have been posted. The problem set key will be posted once I get it cleaned up a bit and I'll have a group NMR activity for you tomorrow.
Many changes have been made to the left: The Need to Knows for Chapters 13, 14 and 15 have been posted, as have the charts for 13C and 1H NMR. (Remember, you'll be given the 13C chart but must learn the 1H chart.) Modeling exercise #10 has also been posted; I'll talk more about it tomorrow.
New changes! A key for the exam and an NMR problem set have been posted. The problem set key will be posted once I get it cleaned up a bit and I'll have a group NMR activity for you tomorrow.
Monday, April 09, 2007
NMR
Today we started NMR spectroscopy. We covered the basics of 13C NMR and were exposed to some basics: upfield and downfield, shielded and deshielded, TMS, ppm, etc. The major point was that every unique carbon atom will have a resonance peak in a 13C NMR spectrum. This means that the most important thing that a student must worry about is determining how many unique carbon atoms are in a compound.
We also looked at two special 13C NMR experiments, DEPT-90 (only shows CH's) and DEPT-135 (shows CH and CH3 peaks pointing up, CH2 peaks pointing down).
We also looked at two special 13C NMR experiments, DEPT-90 (only shows CH's) and DEPT-135 (shows CH and CH3 peaks pointing up, CH2 peaks pointing down).
Thursday, April 05, 2007
IR Spectroscopy
Today we finished our short tour of MS and IR by looking at the basics of infrared spectroscopy. We reduced it down to a half-dozen or so types of bonds that will show up at different portions of the IR spectrum. It is anticipated that you will learn those areas and be able to draw IR spectra of simple molecules, including labels on the axes.
A key to last year's exam has been posted.
A key to last year's exam has been posted.
Monday, April 02, 2007
Spectroscopy

Today we started our treatment of spectroscopy by looking at mass spectroscopy (MS). MS is used by organic chemists primarily as a first look at a compound's molecular weight by observing the molecular ion peak. It is also valuable as a method for determining structure by looking at fragmentation patterns -- a molecule is ionized by bombarding it with an electron beam and the resultant radical cation then fragments into smaller particles. These particles can be detected provided they have a charge. We saw a limited number of fragmentations for which you will be responsible, including alpha cleavages, dehydrations and McLafferty rearrangements.
A new problem set and key has been posted. Last year's exam has also been posted.
The picture is of Asa Larson. He studies the process of an electron recombining with a molecular ion (the reverse of the step that starts MS) in his work at the Royal Institute of Chemistry in Stockholm, Sweden.
Thursday, March 29, 2007
Even More Additions to C=C !!!!

We can't get enough of this addition of various species to alkenes. Yesterday was pretty much the final installment -- addition of H2, ozonolysis and di-hydroxylation were examples of reactions that we just have to know, without mechanistic details. One reaction in which the mechanism is approachable was the radical synthesis of some classes of polymers (we used polystyrene as an example).
A new podcast has been posted. It is a short-form of generic additions of various electrophiles to alkenes.
The image is that of an ozone generator. It is similar to the one that I used to destroy the rubber tubing many years ago.
Tuesday, March 27, 2007
More Alkenes as Nucleophiles
We've continued to look at alkenes as nucleophiles but we have made the electrophiles more complicated than simple HX. Our examples of electrophiles include X2, Hg(OAc)2 and BH3. In those cases, we use available electrons to swing back down to the developing carbocation and eliminate the positive charge on the carbon. In the case of BH3, this is relatively straight forward and takes us to a product in which we have added H-BH2 across a C=C. In the other two cases we make a three-membered ring with a positive charge and we must open the ring by reaction with a nucleophile. Note that the nucleophile must attack the three-membered ring from exactly the opposite side of the leaving group (Br or Hg), so there can be stereochemical implications.
Thursday, March 22, 2007
Alkenes as Lewis Bases/Nucleophiles

Yesterday we looked at how alkenes can react as nucleophiles. The reasoning behind this activity is due to the fact that the pi electrons are not trapped between the nuclei, as are electrons in a sigma bond, but are out in a more open space (remember--side to side overlap of the fat parts of p-orbitals). As a result, the pi electrons can act somewhat like a lone pair of electrons.
The reaction that we looked at was the reaction of alkenes with HBr. It is a two-step process, leading to an intermediate cation which is then attacked by the generated bromide ion. The entire process is just the reverse of an E1 reaction.
The regiochemistry of the reaction is governed by Markovnikov's Rule -- the cation formed is the more stable of the two possibilities. Note that the intermediate cation has an unhybridized p-orbital so the bromide can attack from either side; this can have implications for stereochemistry.
Problem Set 7 and its key have been added to the list of useful links. At this point, we have only covered how to do reaction series (a).
The picture is of Vladimir Markovnikov.
Tuesday, March 20, 2007
Alkenes
Yesterday was a day to start talking about alkenes. We spent most of the time discussing the physical chemistry of the C=C bond -- how it consists of an end-to-end bond and a side-to-side pi bond. I used the example of a pair of forearms and showed how they had to be parallel in order for the bond to exist; as a result it is nearly impossible to rotate around a C=C bond. We also saw the basis for Zaitsev's rule. Essentially, it is hyperconjugation that is responsible for the increased stability of more highly substituted alkenes. At the end, we saw a foreshadowing of the chemistry that we will discuss, the ability of the alkene to act as a Lewis base.
Modeling #7 has been posted; molecule slips will be available tomorrow.
Modeling #7 has been posted; molecule slips will be available tomorrow.
Monday, March 12, 2007
New Postings
I've added the key to exam 2, along with Need to Know links for Chapters 6, 7, 8, and 12. I hope your spring break is off to a great start!
See you next Monday.
See you next Monday.
Monday, March 05, 2007
Work, work, work!
Today we worked pretty hard on being able to determine which reactions are possible for a given set of conditions. All of this is based on looking at the Lewis acid (the compound with the leaving group) and the Lewis base (the nucleophile or base). Once the determination is made which reaction(s) is/are possible, all that remains is to draw the arrows.
We will continue to practice drawing reactions on Wednesday.
We will continue to practice drawing reactions on Wednesday.
Saturday, March 03, 2007
Some Additions to the Links......
Friday, March 02, 2007
Elimination Reactions
Today we covered both the E1 and E2 reactions. In both cases, we use a Lewis base to abstract a proton from the substrate (at a carbon next to the carbon with the leaving group) and form an alkene. There are differences, however: The E2 is a single-step reaction and the stereochemistry of the product is determined at the beginning of the reaction. The E1 reaction is two steps and the presence of the intermediate allows for C-C bond rotation, therefore the stereochemistry is not set until the second step of the reaction.
We started to look at how to determine which of the four possible reactions (SN1,SN2, E1, E2) would take place under a particular set of reaction conditions, with the promise of more to come next Monday and Wednesday.
Over the weekend a copy of last year's exam will be posted, along with a podcast of the material from today's lecture and an elimination problem set. I am also hoping to get some more substitution problems posted to give you more practice with drawing arrows.
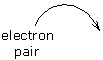
The picture shows what it is all about -- a pair of electrons is moved according to the direction of an arrow, from Lewis base to Lewis acid.
Wednesday, February 28, 2007
SN1!!!
Today we talked about the SN1 reaction, the second of our two substitution processes. There are significant differences between the SN1 and the SN2 reaction: in terms of stereochemistry, an SN1 gives racemization instead of inversion, for steric bulk it is better to have a highly substituted substrate for an SN1 reaction (instead of an accessible carbon like we needed for the SN2), etc. It is important to be able to recognize the difference in energy profiles for the two reactions -- an SN1 is two steps and an SN2 is one step.
There is also a major difference in solvent requirements: in an SN2 reaction, polar protic solvents surround and stabilize the negatively charged nucleophile, slowing the reaction. In an SN1 reaction, you must have a polar protic solvent to stabilize the intermediate carbocation otherwise the energy level is too high and the reaction will not take place.
The podcast for today's lecture has been posted and is available on iTunes. In iTunes, go to Advanced > Subscribe to Podcasts, then enter this URL: http://itech.dickinson.edu/blog/?feed=rss2&cat=765
On Friday I hope to cover both elimination reactions, allowing us to spend next Monday and Wednesday working problems in preparation for Friday's exam.
There is also a major difference in solvent requirements: in an SN2 reaction, polar protic solvents surround and stabilize the negatively charged nucleophile, slowing the reaction. In an SN1 reaction, you must have a polar protic solvent to stabilize the intermediate carbocation otherwise the energy level is too high and the reaction will not take place.
The podcast for today's lecture has been posted and is available on iTunes. In iTunes, go to Advanced > Subscribe to Podcasts, then enter this URL: http://itech.dickinson.edu/blog/?feed=rss2&cat=765
On Friday I hope to cover both elimination reactions, allowing us to spend next Monday and Wednesday working problems in preparation for Friday's exam.
Monday, February 26, 2007
SN2!
Today was a day to learn about SN2 reactions. We pretty much covered it all, from nature of the mechanism to requirements for nucleophiles, leaving groups, substrates, and solvents. A few big points: There is only a single step (no intermediate) and the stereochemistry at the carbon of interest is inverted (the phenomenon is called a Walden inversion). On Wednesday we'll cover the other major substitution reaction, the SN1 (which, confusingly enough, has two steps).
Today's graphics have been assembled into a seven-minute podcast; it will be posted on Tuesday morning and will be available on iTunes.
Today's graphics have been assembled into a seven-minute podcast; it will be posted on Tuesday morning and will be available on iTunes.
Friday, February 23, 2007
Halogens, a rough day

Today was a tough one. We started out by finishing up the treatment of reactions by looking at reaction profiles (or reaction diagrams) and learning some new terminology. The big item: Know the difference between a transition state and an intermediate.
We then covered halogens, both the manufacture (which was heavy on radical chemistry) and one use of them (Grignard reactions). Along the way, we learned a few things about how lower energies for intermediates implies a lower activation energy and a better (faster) reaction, how hyperconjugation stabilizes a radical, and we re-visited allylic stabilization via resonance, but this time on an allylic radical instead of a cation or an anion.
Remember, a brief version of this lecture has been posted on iTunes as "Halides lecture."
The image is of Victor Grignard. An excellent mustache.
Wednesday, February 21, 2007
More on Reactions......
Today started with a group activity in which you had to show the curved arrows for a two-step reaction. This type of problem is central to the subject -- someone learning organic chemistry must learn to draw curved arrows to show the mechanism (electron movement) of a reaction. I showed some strategies for how to solved these problems. Essentially, it boils down to four basic rules: 1) find the Lewis base (anion, lone pair, multiple bond); 2) find the Lewis acid (cation, partial positive charge); 3) draw the curved arrow(s), which must start at the Lewis base and go to the Lewis acid; and 4) don't violate the octet rule! Hopefully, you are able to recognize the relationship between the kinds of arrows that we drew today and those we drew for the standard acid/base reactions that were the subject of the acid/base problem set a few weeks ago.
Monday, February 19, 2007
Exams returned and the start of organic reactions

OK, the return of the first exam was today, with a brief run-through to show where most of the problems seemed to occur. Check the key, come to me with questions if necessary. If you suddenly find that you are not performing up to your standards/expectations, you need to make sure that you are getting help -- either me, the group study sessions, or a tutor.
Today we started looking at Chapter 5, which deals with reactions. The main point was to show that we can look at reactive processes using the same methods as we used for resonance structures and acid/base reactions -- curved arrows, indicating electron movement.
We saw two kinds of electron movement in bond cleavages -- homolytic (both sides of a bond are treated the same way and each atom receives an electron; use "fish hook" arrows) and heterolytic (the sides are treated differently and one atom receives both electrons and the other is given none). The direction that the electrons flow in a heterolytic cleavage is governed by the polarity of the bond -- the electrons flow in the direction of the atom that is happier to have excess electrons. Generically, this is decided by electronegativity. The concept is made a bit simpler by the realization that nearly all heteroatoms (anything other than a C or H) attached to a carbon is more electronegative than the carbon and will grab the electrons from a carbon-heteroatom bond, leaving C with only six electrons and a more positive charge.
The picture is of a "rotating bomb combustion calorimeter," an instrument used to measure the energy contained within a compound. In this particular picture, it is being used to measure the differences between fuel oils containing differing amounts of heteroatom additives.
Friday, February 16, 2007
Diastereomers and Meso compounds

Today we started off with some practice at identifying (R) and (S) configurations -- and saw that there is still some work to be done. Remember, the substituent of priority number 4 MUST be going back for the CIP system to work. Futhermore, if you switch two substituents to get to that scenario, the act of switching has changed the stereochemistry to give you the opposite of the stereochemistry of the original compound.
We also looked at scenarios with more than one stereocenter and saw that there are a few possibilities that can come about. If every stereocenter is changed, then you have made the enantiomer UNLESS there is a plane of symmetry in the molecule. In that case, it is a meso compound and the mirror images are identical. If at least one but not all of the stereocenters are changed, then you have made a diastereomer of the original compound; the two compounds most likely have different physical attributes.
The image shows the addition of dioxygen to a compound to make a pair of diastereomers -- one is R, S, S, R and the other is R, R, R, R. The slash through the lower arrow shows that the pathway to the R, R, R, R compound does not happen.
Thursday, February 15, 2007
Stereocenters
Yesterday was a day to learn about stereocenters. In organic chemistry, that essentially means a carbon with four different substituents attached to it. We was in class that such a carbon is not superimposable upon its mirror image -- we termed these things enantiomers. We also learned how to use the R and S descriptors for identifying the stereochemistry at a carbon center.
Subscribe to:
Posts (Atom)